Machine learning potentials for modeling alloys across compositions - Science | AAAS
This story deserves to be shared
Send it to someone who'd care, or post it on your networks.
This article is an excerpt. To read the full content, visit the original source. Science | AAAS
Related articles
Synthesizability, hardness, and stacking order in multicomponent transition metal carbides from machine-learned potentials - Nature
We’re sharing this article early to provide faster access to peer-reviewed, accepted research. It is citable and carries a permanent DOI. This version is subject to further edits and will be replaced automatically by the final Version of Record. All legal disclaimers apply. AbstractMulticomponent transition metal carbides are promising for extreme-environment applications, but identifying compositions that are both synthesizable and hard remains challenging. We fine-tune the MACE machine-learned interatomic potential on approximately 28,000 density functional theory calculations spanning the composition space of groups 4–6 transition metals and carbon to predict the thermodynamic stability and elastic properties of multicomponent carbides. The fine-tuned model achieves formation energy errors of ~10 meV/atom for thermodynamically relevant structures with only 20% of the training data. We screen over 1500 equiatomic compositions across rocksalt, hexagonal, and hcp prototypes, combining free energy models with elasticity-based hardness surrogates. Synthesizability predictions at 1500 °C agree well with experimental reports for both single-phase and multiphase carbides.
HIP: Hessian Interatomic Potentials without derivatives - Nature
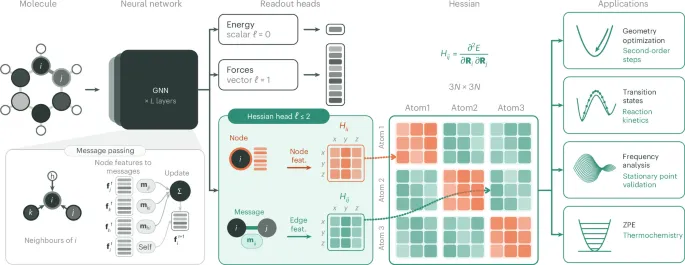
MainMolecular simulation is a cornerstone of material discovery and molecular design. While the prediction of energy and forces is often sufficient for structural dynamics, advanced computational chemistry workflows often depend on the molecular Hessian1,2. By describing the local curvature of the potential energy surface (PES), the Hessian is foundational to many computational workflows. Second-order geometry optimization relies on the curvature information to rapidly and reliably converge to equilibrium structures3. Transition state searches and extrema classification depend on the Hessian to correctly identify saddle points and reaction barriers, which are essential for parameterizing reaction rates and microkinetic models for longer time- and length-scale simulations4. Furthermore, mode-following transition state search methods directly leverage the curvature information to locate the transition states5. Finally, vibrational analysis uses Hessian eigenvalues to compute zero-point energy (ZPE) corrections and thermodynamic free energies bridging the gap from static molecules to nonzero temperature observables6.
Machine learning modeling and interpretation of saline soil thermal conductivity - Nature
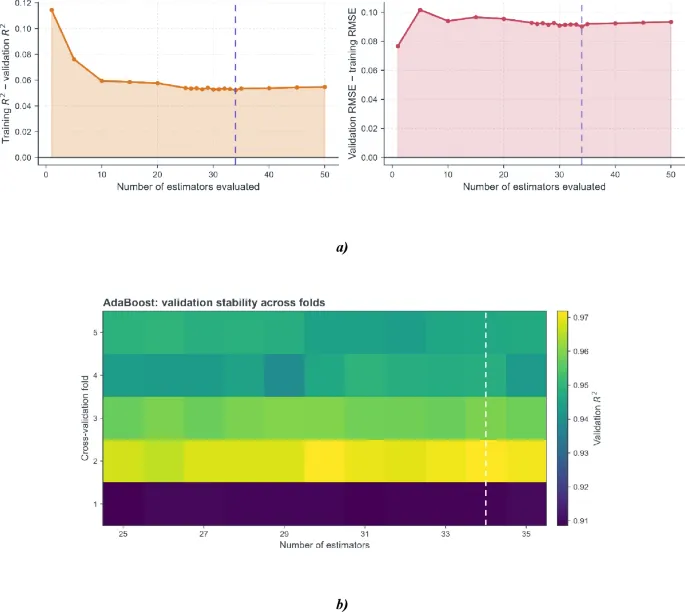
We’re sharing this article early to provide faster access to peer-reviewed, accepted research. It is citable and carries a permanent DOI. This version is subject to further edits and will be replaced automatically by the final Version of Record. All legal disclaimers apply. AbstractSoil thermal conductivity is a key parameter in geothermal design, buried infrastructure, frozen-ground engineering, agricultural heat-transfer assessment, and subsurface energy systems. This study developed an interpretable multi-model machine-learning framework for predicting soil thermal conductivity from 903 literature-derived observations and 15 predictors describing temperature, texture, bulk density, volumetric water content, salt concentration, salt molar mass, and ionic identity. Eleven regression algorithms were optimized by five-fold cross-validation within a common 90% development subset and compared on an untouched 10% random hold-out. Extra Trees yielded the lowest observed random-holdout error (R² = 0.9816, RMSE = 0.0625 W m⁻¹ K⁻¹, MAE = 0.0446 W m⁻¹ K⁻¹, AARE = 3.93%, and A20 = 98.90%), with CatBoost and XGBoost providing similarly high point performance.
Zoya Abbasi and Jaden Saïd win 2026 Applied Mathematics, Modeling and Computational Science (AMMCS) prize - University of Waterloo
Congratulations to Zoya Abbasi and Jaden Saïd, winners of the 2026 Applied Mathematics, Modeling and Computational Science (AMMCS) prizes at this year’s AMMCS International conference. AMMCS aims to promote interdisciplinary research and collaboration involving mathematical and computational sciences within a larger international community, and highlight recent advances in applied mathematics, modeling and computational science. The interdisciplinary international conference started in 2011 and was hosted in Waterloo this year.
How Claude is uplifting biomolecular modeling - Anthropic
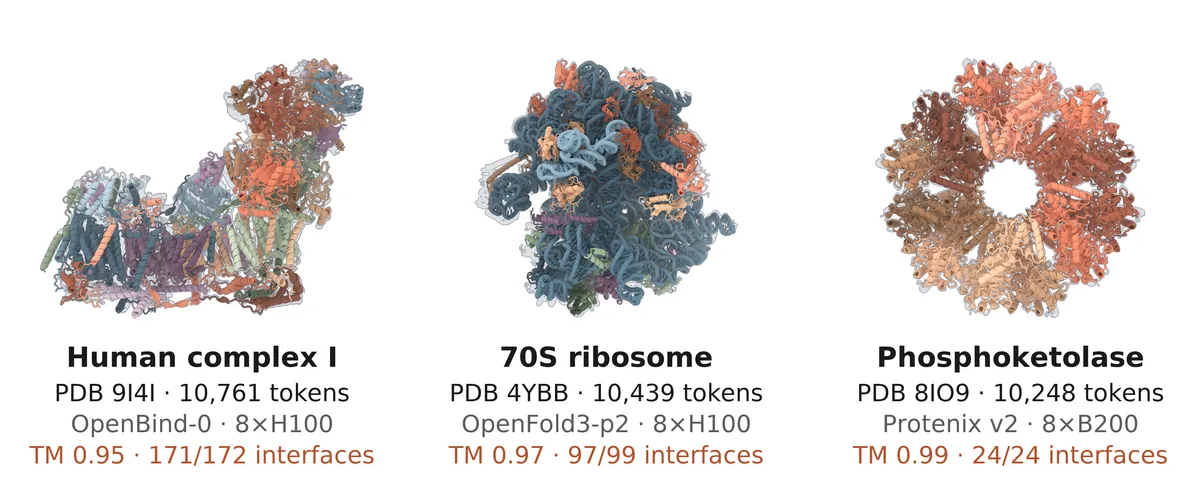
In this post, we share how Claude made the open-source models that scientists use to predict and design biomolecules faster and more memory-efficient. Claude, working within Claude Science, optimized more than 30 of these models in just under four weeks, speeding them up roughly 4x on average. It also created a low-memory mode that enables the accurate prediction of biomolecular systems larger than 10,000 tokens (amino acids, nucleotides, and atoms from small molecules and ions) on a single NVIDIA GPU node. We are open-sourcing all of the optimized code and announcing a protein design competition co-sponsored with Adaptyv Bio, backed by up to $1 million in Claude credits and wet lab validation for over 5,000 designs. Recently, we shared results demonstrating Claude’s abilities to design de novo protein binders through expert-level orchestration of open-source protein design and structure prediction models. De novo binders are small, computationally designed proteins that attach tightly to a specific target molecule to activate, block, or deliver something to it.
0 comments
No comments yet. Be the first to share your thoughts!